Dossier assembly and review · Training guide
IND & IMPD Quality
Write the quality part of an IND or an EU investigational medicinal product dossier (IMPD), appropriate to the phase: drug substance and drug product summaries, impurities compared with the toxicology batch, the specification, and a shelf life that covers the trial.
Open the tool at https://app.cmcfy.com/app/ind-impd-quality.html. This guide is also online at https://cmcfy.com/guides/ind-impd-quality.html
1. What this module does
You enter the phase, dosage form, maximum clinical dose and trial duration, the impurity levels in the toxicology batch and the clinical batch, the specification and the stability data.
The tool qualifies each impurity: covered by the toxicology batch, covered by the daily intake at the NOAEL, or below the ICH Q3A threshold. It checks the tests expected for the phase, form and product type, the EU shelf-life extrapolation rule, and that the shelf life covers the trial.
What you get
- US: IND 312.23(a)(7) drug substance and drug product sections.
- EU: IMPD 2.1.S and 2.1.P quality sections, with placebo where supplied.
- A Word or PDF file for each.
2. Before you start
Have these ready. You can still start with the example data and replace it piece by piece.
- Phase, dosage form, maximum daily dose and trial duration including supply.
- Toxicology batch number and NOAEL.
- Impurity levels in the toxicology and clinical batches, with mutagenic alerts.
- Composition, specification, method suitability, stability data and extension plan.
3. Step by step with the example
Open the tool at app.cmcfy.com/app/ind-impd-quality.html. The example is loaded when the page opens, so every panel already has realistic data. Follow the steps to see how it works, then replace the example with your own data.
Describe the trial
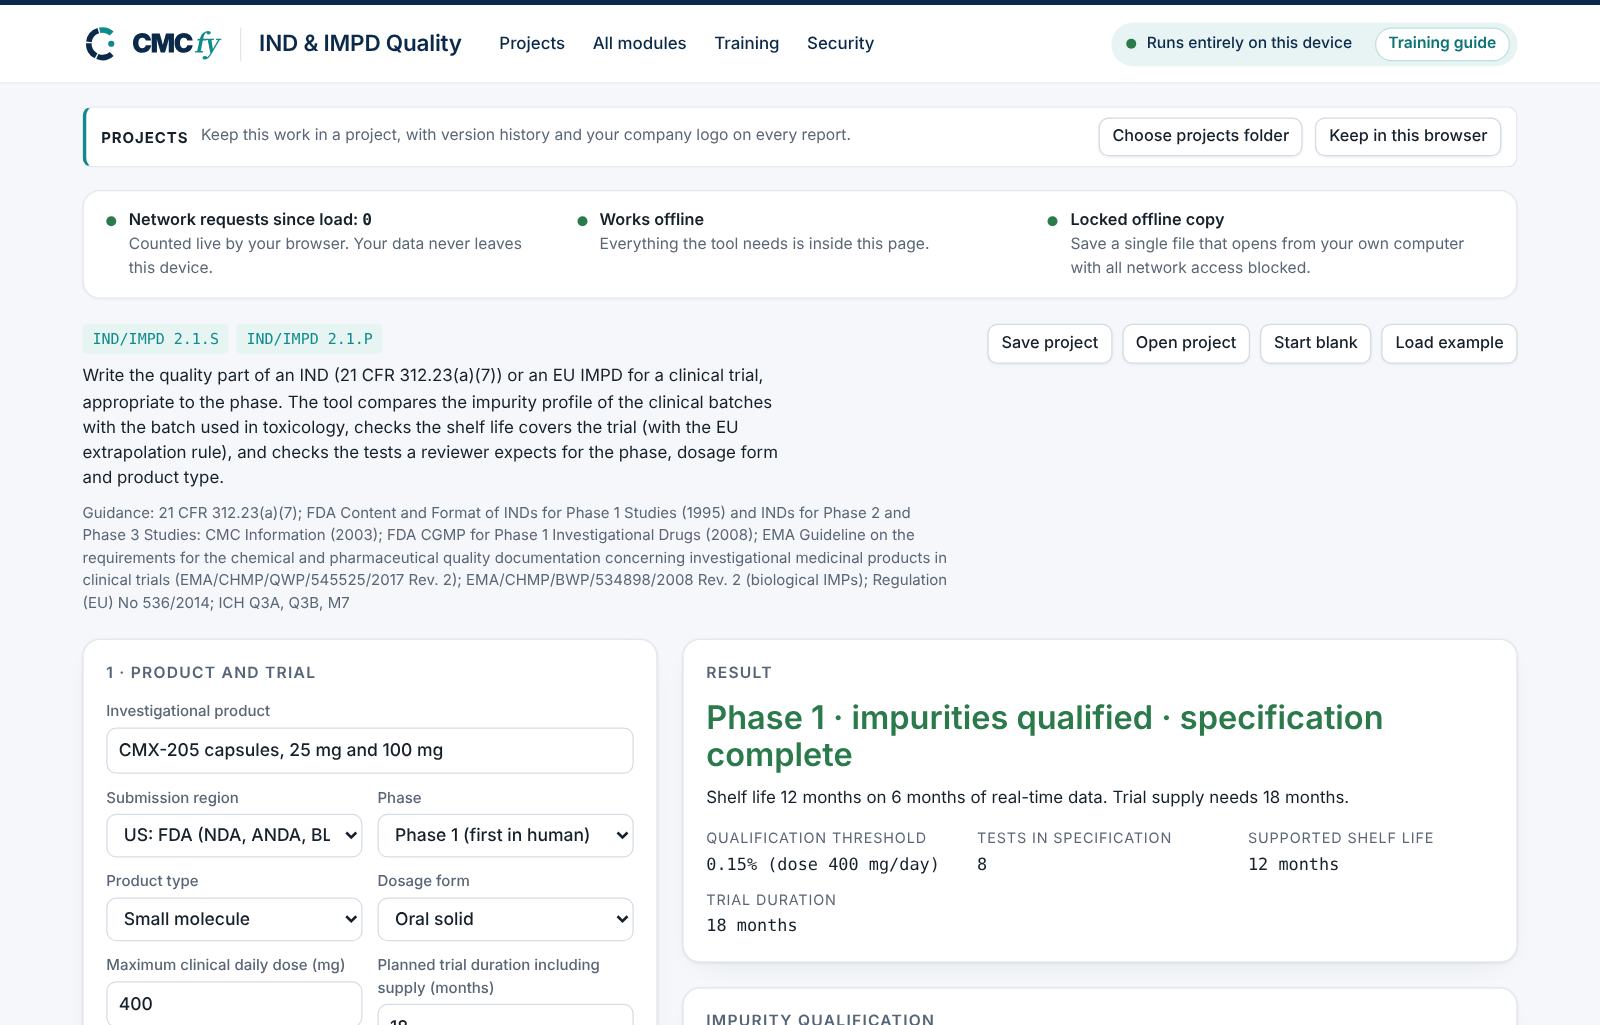
Choose the region, phase, product type and dosage form, and enter the maximum daily dose and the trial duration.
Enter impurities and the specification
One row per impurity with the level in the toxicology batch and in the clinical batch, the clinical batch number, and yes if it has a mutagenic alert. One row per test in the specification.
Read the result
The headline gives the phase, whether every impurity is qualified and whether the specification has the expected tests: "Phase 1 · impurities qualified · specification complete".
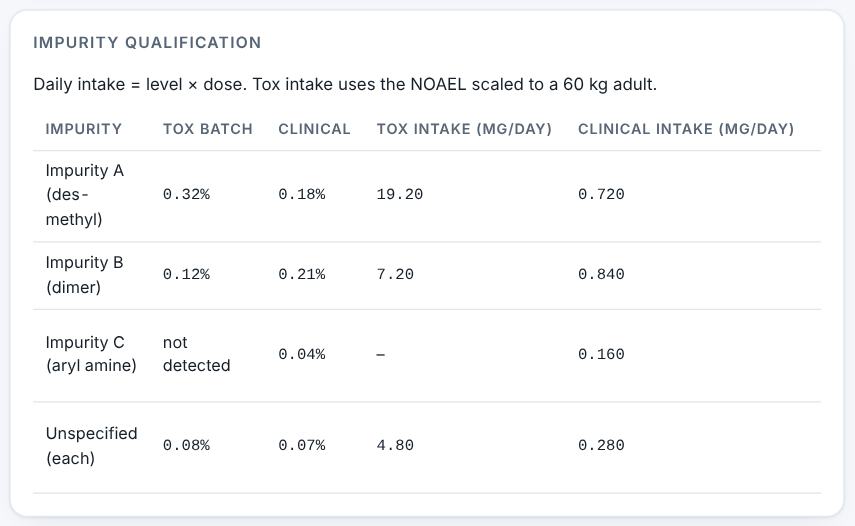
Impurity qualification. Resolve alerts and download
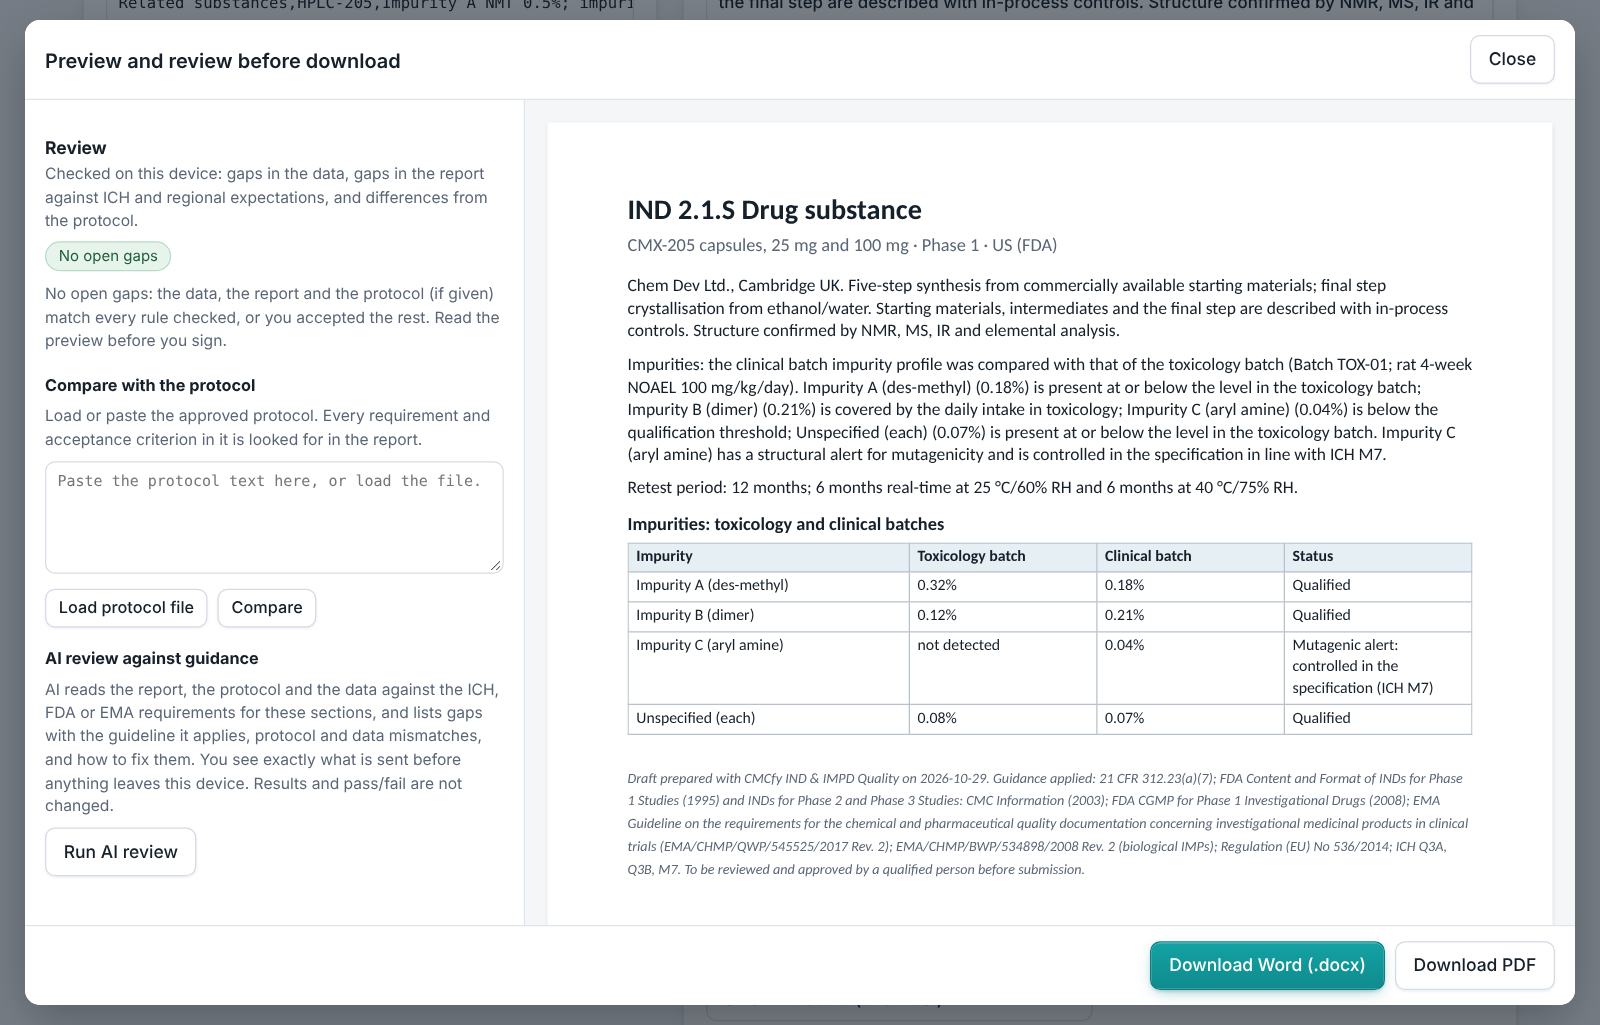
Unqualified impurities, uncontrolled mutagenic impurities, missing tests and a shelf life beyond the EU rule are flagged. Then preview and download.
4. Field reference
Every input in the tool, panel by panel. Tables accept rows pasted straight from Excel, comma-separated text, or a loaded file; check the columns after loading a file. Fields marked “when it applies” appear only for the choices that need them.
1 · Product and trial
| Field | What to enter |
|---|---|
| Investigational product | Type it in. |
| Submission region | Choose one: US: FDA (NDA, ANDA, BLA, IND); EU: EMA and national agencies (MAA); Both US and EU. |
| Phase | Choose one: Phase 1 (first in human); Phase 2; Phase 3. |
| Product type | Choose one: Small molecule; Biological. |
| Dosage form | Choose one: Oral solid; Oral liquid; Parenteral (sterile); Other. |
| Maximum clinical daily dose (mg) | Type a number. |
| Planned trial duration including supply (months) | Type a number. |
| Placebo or comparator supplied | Tick the box if it applies. |
2 · Drug substance
| Field | What to enter |
|---|---|
| Manufacturer, route and controls | Type or paste text. |
| Drug substance retest period and real-time data (months) | Type it in. |
3 · Impurities: toxicology batch against clinical batches
Levels in % of the drug substance. The tox dose is the NOAEL dose in the most sensitive species in mg/kg/day; human dose uses 60 kg.
| Field | What to enter |
|---|---|
| Toxicology batch and NOAEL | Type it in. |
| NOAEL (mg/kg/day) | From the pivotal toxicology study in the most sensitive species. Type a number. |
| Impurities | Leave a level empty if the impurity was not detected in that batch. A table. Columns: Impurity | Toxicology batch (%) | Clinical batch (%) | Clinical batch | Mutagenic alert (yes/no). Paste rows from Excel, type comma-separated rows, or press Load file (Excel, CSV, Word, PDF or a scan). |
4 · Drug product
| Field | What to enter |
|---|---|
| Composition and manufacture | Type or paste text. |
| Specification | A table. Columns: Test | Method | Acceptance criterion. Paste rows from Excel, type comma-separated rows, or press Load file (Excel, CSV, Word, PDF or a scan). |
| Placebo when it applies | Type or paste text. |
| Method suitability | Type it in. |
| Proposed shelf life (months) | Type a number. |
| Real-time stability data on the clinical batch or a representative batch (months) | Real-time months on the clinical batch or a representative batch. Type a number. |
| Stability programme and extension plan | Type or paste text. |
5. Checks and alerts
Calculations and pass/fail outcomes always come from fixed rules, never from AI. The tool checks:
- Qualification threshold min(0.15%, 1.0 mg/day) (0.05% above 2 g/day). Tox intake = level × NOAEL × 60 kg; clinical intake = level × dose.
- Mutagenic impurities named in the specification count as controlled per ICH M7.
- Expected tests for the phase, dosage form and product type (for example sterility and endotoxin for parenterals, potency for biologics).
- EU: shelf life no more than twice the real-time data and no more than 12 months beyond it.
How to resolve alerts
- Critical alerts block the download and must be fixed in the data. A failing result can instead be linked to an investigation: type the investigation reference and outcome and press Record.
- Major alerts block the download until you fix them or type a justification and press Accept justification.
- Minor alerts do not block anything. They are listed in the documents so the reviewer sees them.
- Some alerts have an Apply fix button that makes the change for you. Reopen undoes a justification.
Narratives
This module writes: Drug substance summary; Drug product summary. Each narrative is written by fixed rules from your data. You can edit it in the box. Rewrite with AI (where your plan includes AI) rewrites it in regulatory style; every number in the text is checked against your data, and a number that does not match is flagged under the box and blocks the download until you correct it. Reset to rule-based text brings back the original.
6. Preview, gap review and download
- Press Preview next to a section, or Preview and download all. The document opens in a preview window, exactly as the Word file will read.
- The left side lists the review, run on your device: gaps in the data (missing values, open alerts), gaps in the report (missing content the guidance expects, placeholders such as [site] or TBD left in the text) and, if you load the approved protocol under Compare with the protocol, differences from the protocol (requirements, criteria, batches or time points it plans that the report does not show).
- Click a finding to jump to the place in the preview. Fix it in the tool, then preview again.
- Optional AI review: a second read for contradictions, claims without data and vague wording. The report text is sent to Claude only when you press it, and only where your plan includes AI. Check every AI finding; AI can be wrong.
- Press Download Word (.docx) or Download PDF. With several documents open, All as Word (.zip) and All as PDF (.zip) download the set. The download buttons stay disabled while a blocking alert is open; the preview still works.
Documents this module produces
| Section | Document |
|---|---|
| 2.1.S / IND DS | Drug substance summary, impurity comparison |
| 2.1.P / IND DP | Composition, specification, stability, placebo |
Use Save project to keep your work as a file on your computer, and Open project to continue later. Start blank clears every field; Load example brings the example back. Nothing is stored on CMCfy servers.
7. US and EU submissions
US follows 21 CFR 312.23(a)(7) and the FDA guidances on content and format of Phase 1 INDs (1995) and Phase 2 and 3 CMC information (2003). EU follows EMA/CHMP/QWP/545525/2017 Rev. 2 (chemical IMPs) or EMA/CHMP/BWP/534898/2008 Rev. 2 (biological IMPs) under Regulation (EU) No 536/2014.
8. Working with other modules
- Stability data come from the Stability Studio.
- At marketing application, write the full Module 3 sections with the QOS Composer & Dossier Assembler.
9. Common pitfalls
- Comparing percentages without the dose: an impurity at a higher percentage can still be covered by the daily intake in toxicology.
- Proposing a shelf life longer than the EU rule allows without a stability commitment.
10. Practice exercises
Each exercise starts from the example (press Load example to reset). We run these on every update of this guide, so the result you see should match.
Lower the NOAEL
Set NOAEL (mg/kg/day) to 5.
Expected resultThe headline shows one impurity not qualified.
Extend the shelf life in the EU
Set the region to EU and Proposed shelf life to 18.
Expected resultThe note says the EU rule supports up to 12 months.